Regulatory Classification of Nanoparticle Drug Products at the FDA
The FDA uses existing drug and biologic frameworks to evaluate nanoparticle products.
Nanoparticle nose-to-brain delivery for CNS peptide indications must fit the FDA’s existing drug or biologic frameworks. Nanocarriers are designed to use trigeminal and olfactory pathways to reach the brain, but that delivery concept does not determine regulatory classification. The practical strategy is to establish the product’s regulatory identity early, then build evidence that connects its nanoscale properties, manufacturing controls, and intended route of administration.
Why the FDA has no nanomedicine category and what it uses instead
Developers looking for a “nano lane” encounter a basic problem before they reach the details of a submission. The FDA has not established a regulatory definition for “nanotechnology,” “nanomaterial,” or related terms. Instead, the agency evaluates the product through existing processes, with safety, effectiveness, and regulatory status addressed individually.
The FDA’s April 2022 final Guidance for Drug Products, Including Biological Products, that Contain Nanomaterials describes how this works. The agency’s position leaves room for products with very different active ingredients and carrier systems to be assessed under the same broad standards. It also means that the presence of nanoparticles is a reason to examine their role.
The EMA also assesses nanotechnology-based medicines under existing pharmaceutical and biological product frameworks. The comparison matters because the lack of a separate category is not simply a naming choice unique to the FDA. Both agencies use existing regulatory structures for new delivery systems, though those structures do not fully account for nanoscale-dependent behavior. This creates two distinct questions for a developer, namely where the product belongs and what evidence its unusual properties require. Resolving the first does not resolve the second.
How the existing drug, biologic, and combination-product frameworks absorb nanoparticle products
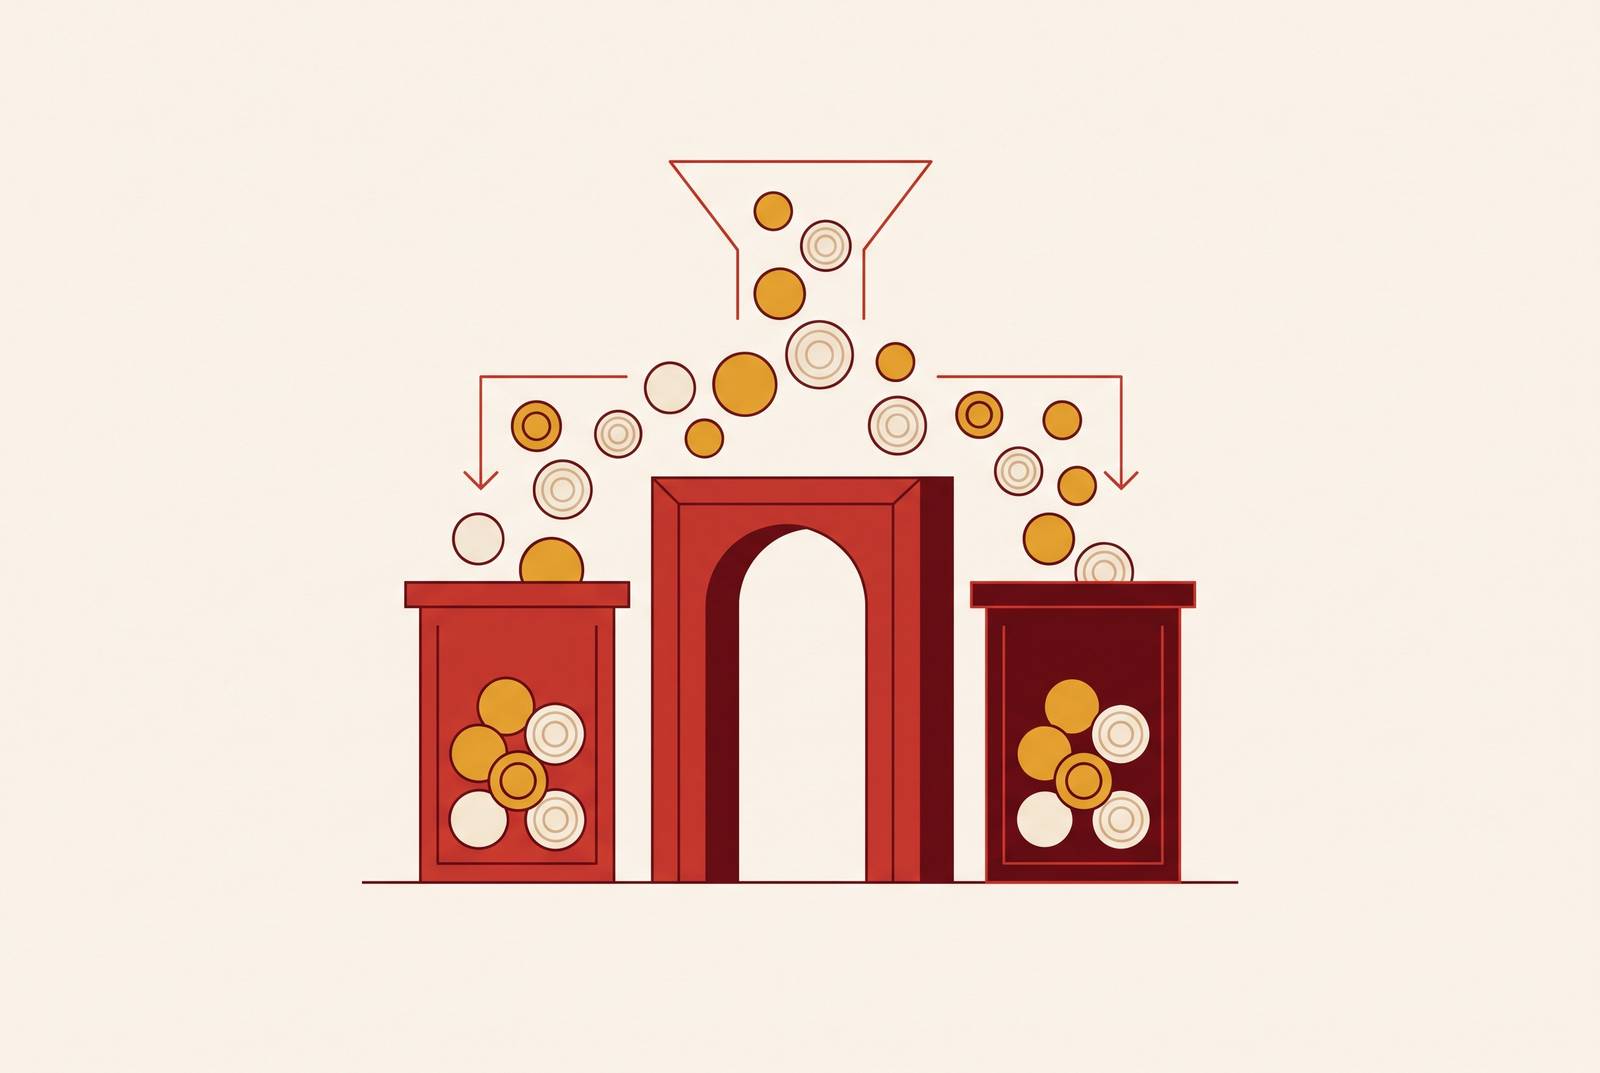
Primary mode of action is the starting point for this map. Chemical action points toward classification as a drug, mechanical action toward classification as a device, and biological action toward classification as a biologic. Nanoparticle products fall into these existing categories and are assessed one by one as complex products. The carrier’s dimensions do not replace the analysis of what the product does. When drug, device, and biologic functions overlap, combination-product classification procedures offer a way to assign a lead center.
The nature of the active ingredient makes this map more specific. Nanoparticles delivering small molecules fall within the drug framework through the NDA/ANDA route. Products delivering large proteins or certain nucleic acids such as mRNA may fall within the biologic framework through the BLA route. However, short peptides of ≤40 amino acids and some nucleic acid therapeutics such as siRNA are regulated as drugs through the NDA route. These distinctions make “peptide nanoparticle” an insufficient description for assigning a pathway.
For peptide-delivery systems such as Lionbio’s intranasal GLP-1 platform, the classification analysis therefore turns on the active principle. A biologic classification would bring the product into the BLA framework, while the short-peptide distinction can point toward a drug submission. The intended biological effect alone does not remove the need to establish which category applies to the particular peptide and formulation.
Approved nanoformulations of paclitaxel and doxorubicin show how conventional classification operates in practice. They proceeded as drugs rather than through a new nanoparticle category, while liposomes and protein-based formulations have proceeded under drug or biologic classifications. A different example is CNM-Au8, oral gold nanocrystals pursuing an NDA submission in Q3/Q4 2026 for ALS under the accelerated approval pathway, based on neurofilament light chain biomarker data. That planned submission illustrates that even an inorganic nanoparticle therapeutic can pursue a conventional regulatory route. Developers can use these routes, though the scientific work each requires can be hard.
Why nanoscale properties create CMC burdens that conventional frameworks were not designed to handle
Once classification is settled, the central difficulty shifts to chemistry, manufacturing, and controls, or CMC. A submission must characterize and control properties that can behave differently at the nanoscale than they do in bulk materials under the same manufacturing conditions. The critical quality attributes, or CQAs, include particle size, size distribution (PDI), surface charge, composition, and drug loading efficiency. Each can be highly sensitive to manufacturing conditions in ways that small-molecule CQAs typically are not.
The harder question is how those measurements connect to performance. There is no thorough understanding of the relationship between manufacturing process parameters, CQAs, and in vivo clinical performance across these systems. Consequently, a change in process or scale cannot reliably be translated into a prediction of therapeutic behavior. Measuring particle size is not the same task as establishing what a size change means for a particular product.
Conventional regulatory expectations do not fully account for size-driven biodistribution, dynamic surface interactions including bio-corona formation, or sensitivity to manufacturing variation. Batch-to-batch variability is a principal obstacle for the field, together with bio-corona formation and organ accumulation concerns. For peptide and biologic nanoformulations, limitations in standardization and production add to the burden. These are engineering and scientific problems that a conventional pathway does not eliminate.
The FDA’s CDER perspective does not assume that a product is harmful or benign because it contains a nanomaterial. It applies the same standards of safety, efficacy, and quality, while making characterization and the intended role of the nanomaterial central to evaluation. A risk-based approach also governs the discriminating in-vitro release method. The task is therefore not to argue that a nanoparticle deserves different standards, but to establish how the product can meet those standards with evidence suited to its properties. Rigorous characterization makes the gap navigable without pretending that every process-to-performance relationship is already understood.
How does nanoparticle nose-to-brain delivery for CNS peptide indications compound classification decisions
With brain targeting, the evidence needed also depends on the route, the mechanism of CNS entry, and the neurological target. The blood-brain barrier restricts systemic delivery of GLP-1 receptor agonists, making access to the intended tissue a first-order scientific and regulatory issue. For CNS-directed development, the relevant questions are which framework covers the active ingredient, how the delivery system is intended to reach the brain and which properties govern that process. Classification remains necessary, but it cannot stand in for a route-specific account of the product.
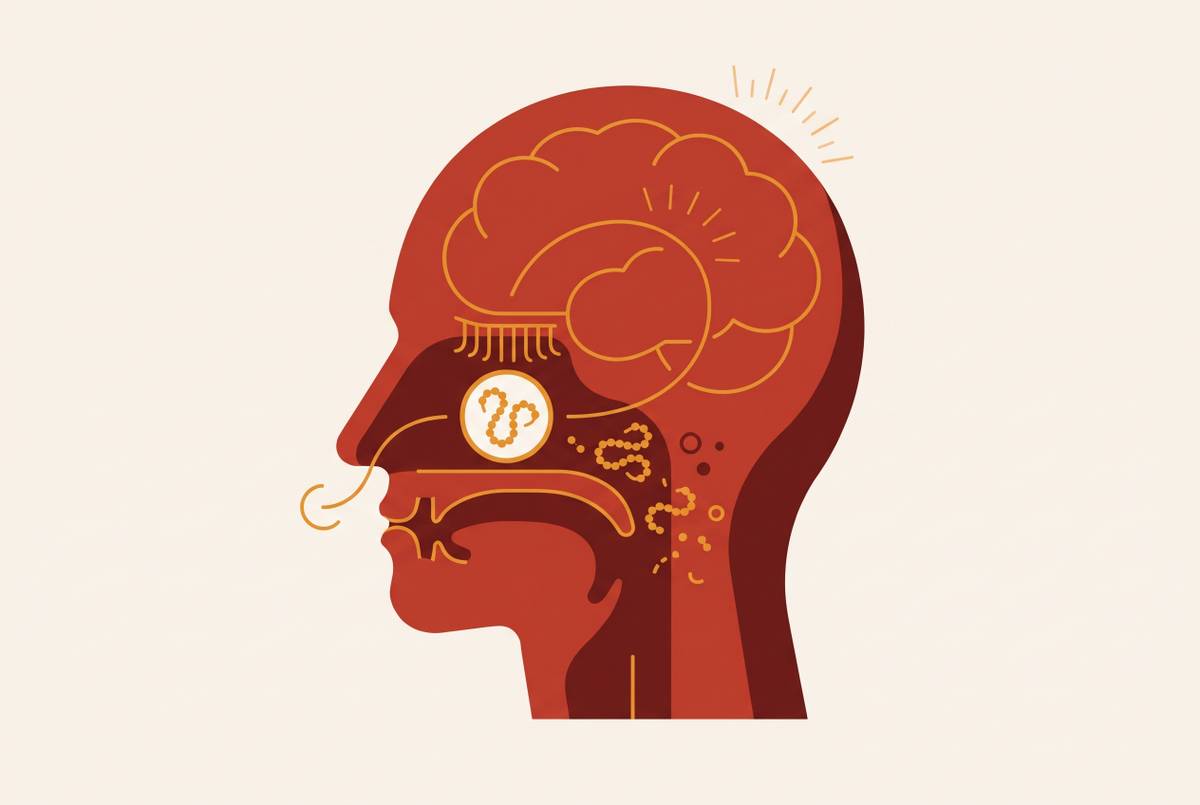
Nose-to-brain delivery could offer a way around the blood-brain barrier through the trigeminal and olfactory pathways. The trigeminal pathway is the dominant route for intact nanoparticles, whereas the olfactory pathway is more likely to deliver substances already released from nanoparticles. This distinction separates carrier transport from payload transport. It therefore has direct implications for which CQAs must be controlled and documented. Evidence about the carrier cannot automatically answer every question about the material released from it.
The nasal route also requires developers to address enzymatic degradation, mucociliary clearance, and limited residence time on the nasal mucosa. Nanocarrier engineering must address these constraints through systems including lipid-based, polymeric, protein-based, or inorganic carriers intended to protect the payload and extend residence time. For peptide formulations, surface interactions, size-driven distribution to specific CNS compartments, and manufacturing sensitivity deepen the characterization burden. Traditional peptide CMC expectations alone do not capture that full intersection. The route is part of the product’s intended mechanism.
Lionbio is developing an intranasal nanoparticle platform designed to deliver GLP-1 peptides through the nose-to-brain route. In this program, classification and nasal-route CQAs need to be considered together from the earliest IND-enabling studies. The relevant submission could be a drug or biologic submission depending on the peptide and formulation.
Preclinical findings support feasibility while leaving substantial translational questions open. GLP-1 and GLP-2 derivatives with functional sequences were rapidly transported through trigeminal axons after intranasal application in mice. King’s College London used gold nanorods with high therapeutic loading capacity for intranasal acyl-ghrelin delivery in a 2025 study, building on its previous work showing that intranasal gold nanorods reached the brain with minimal systemic exposure. These findings do not establish clinical effectiveness for another formulation or peptide. Comprehensive clinical investigations remain necessary, and no approved product has yet fully traversed the regulatory pathway for intranasal nanoparticle CNS delivery.
What the absence of harmonized nanoparticle standards costs developers in practice
Case-by-case evaluation without harmonized nanomedicine-specific standards creates uncertainty about regulatory expectations and approval timelines. Developers face the usual challenge of generating persuasive evidence, but also uncertainty about how nanoscale-specific evidence will be assessed. That distinction matters because it can increase development cost and time even after the broad submission pathway is known.
For peptide nanoformulations, the lack of clearer guidance addressing nano-like biologics makes this problem particularly concrete. Two developers with scientifically similar products may encounter different data requests, review standards, and timelines depending on reviewers and precedents. This does not mean the products should receive identical decisions or that individual review is inappropriate. The structural cost is uncertainty over the work needed to make a product’s case, in addition to uncertainty over whether that case will succeed.
The small approved product base, despite decades of preclinical promise and a growing clinical trial footprint, underscores the broader translational gap. Regulatory uncertainty contributes to that gap, but it is not a complete explanation for it. Manufacturing variability and incomplete understanding of nanoscale behavior remain substantive obstacles in their own right. Industry still seeks clarity on some points after the April 2022 final guidance, which followed the 2017 draft.
What a practical regulatory strategy looks like for nanoparticle therapeutics given the current framework
A defensible strategy begins with an early, documented classification decision and proactive engagement with FDA reviewers before the IND. Primary mode of action and the active ingredient determine the submission framework, while nanoscale properties determine much of the characterization work within it. For a peptide product, that analysis must explicitly account for the distinction between drug and biologic classification. Where the BLA framework applies, characterization must address both the peptide and the nanoparticle carrier. Treating the carrier as incidental would miss the central scientific reason that the product requires deeper scrutiny.
Where a product spans drug, device, and biologic functions, a formal assignment mechanism can reduce ambiguity before major development investments. The RFD process is codified in 21 CFR Part 3, and OCP was established in 2002. These combination-product procedures offer a way to request FDA assignment of a lead center. Early talks with the agency allow developers to plan the evidence as they work through classification. Continuous discussion is especially important where the product raises questions that available precedents do not settle.
For intranasal CNS products, the CMC package must cover nasal-route CQAs as well as standard nanoparticle attributes. Enzymatic degradation protection, mucoadhesion, and particle integrity at the mucosal surface belong beside size, PDI, surface charge, and drug loading. The aim is a documented scientific account of how the formulation supports its intended delivery mechanism. That account must also explain how sensitive the product is to manufacturing variation.
AI-assisted manufacturing controls and frameworks that build safety into design may in time reduce case-by-case variability, but programs must now work within the framework in place. The immediate priorities are to understand classification logic, control CQAs rigorously, and engage the agency early and continuously. The regulatory demands are substantial because the science is genuinely novel. Developers that invest in characterization depth and early engagement are best positioned to turn that complexity into a durable competitive advantage.
Sources
- Nanomedicine in 2026: Illustrative Quantitative Analyses of EPR Heterogeneity, Clinical Trial Attrition, and Emerging Horizons for Active Nanotherapeutics - PMC
- Regulatory framework for polymer-based nanotherapeutics in clinical translation - PMC
- Nanoparticle Therapeutics in Clinical Perspective: Classification, Marketed Products, and Regulatory Landscape
- Nanoparticles in Clinical Trials: Analysis of Clinical Trials, FDA Approvals and Use for COVID-19 Vaccines
- Current state of nanomedicine drug products: An industry perspective - ScienceDirect
- Nonclinical Perspective on Development of Drug Products Containing