Nanoparticle Scale-Up From Lab Bench to GMP Manufacturing
Controlling particle physics, not just batch volume, keeps nanoparticles consistent at scale.
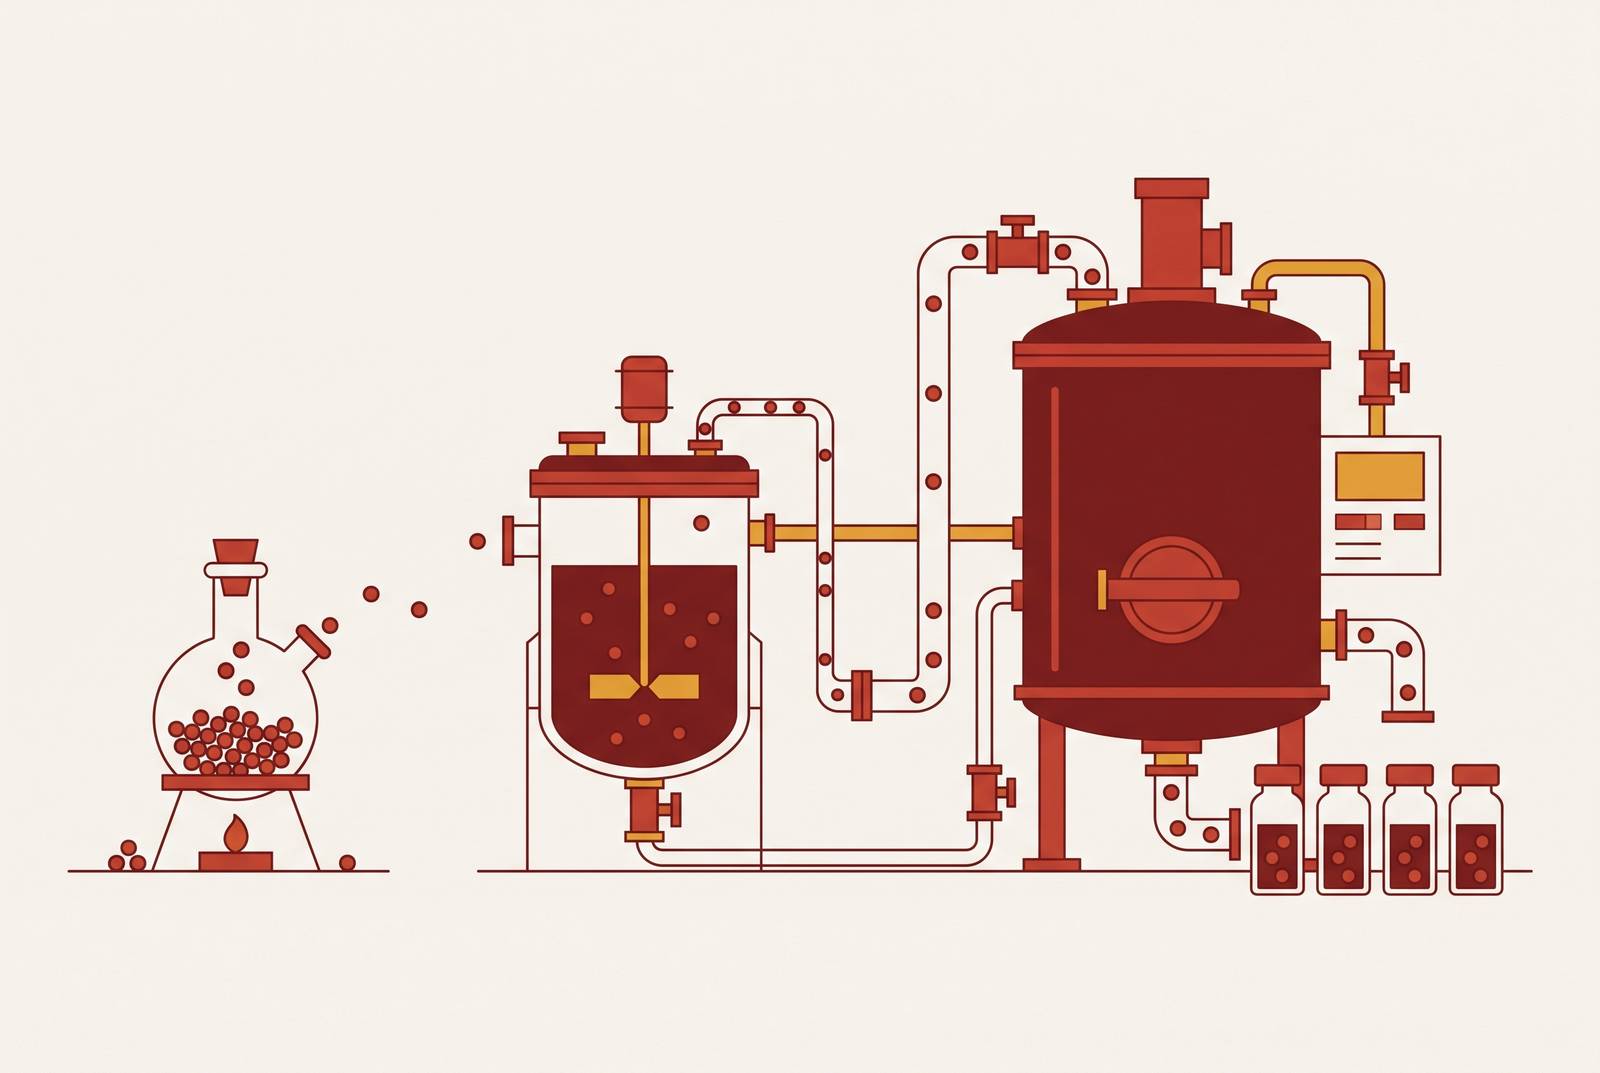
Scaling nanoparticles from lab bench to GMP manufacturing means preserving the physics that defines the product while increasing production. For nanoparticle nose-to-brain delivery for neurological peptide indications, that means controlling both particle formation and the formulation properties needed for olfactory delivery. Intranasal platforms aim to deliver peptides directly to brain pathways, but this depends on keeping the formulation’s properties intact in the material made for clinical use.
Why nanoparticle scale-up is a physics problem, not a volume problem
A bigger batch of salt tablets is still salt tablets, at least in the intuitive picture of pharmaceutical manufacturing. Conventional tablets benefit from decades of industrial experience connecting batch size with product quality. Nanoparticles have no equivalent heritage, and the analogy breaks at particle formation. Their properties depend on both the ingredients in the recipe and the physical environment created by the equipment.
A change in equipment can thus change the product even when the formulation stays the same. Mixing regime, flow dynamics, and shear forces govern how particles form and what size distribution they have. A shift in those conditions can invalidate development work or produce an out-of-specification batch that cannot enter clinical trials.
The critical process variables in microfluidic LNP manufacturing and what happens to them at scale
For lipid nanoparticles made by microfluidics, three critical variables are total flow rate, flow rate ratio (FRR), and mixing geometry. FRR is the primary determinant of initial particle size, with a higher aqueous-to-organic ratio producing smaller particles. Holding that ratio constant across scale transitions preserves a key condition of particle formation. Increasing throughput is a separate task that calls for parallel channels or larger devices, rather than just a higher pump speed on the equipment already in use.
Higher total flow rates improve mixing efficiency but also increase shear stress on the particles. Both effects must be measured at each scale, since better mixing alone does not ensure a better product. Mixing geometry adds another constraint because different microfluidic chips produce different mixing regimes. A larger chip is not automatically equivalent to the development chip, even when both come from the same manufacturer.
Lipid concentration in the organic phase and payload concentration in the aqueous phase affect encapsulation efficiency through their relative proportions, independently of FRR. Keeping the flow ratio the same is not enough to show that encapsulation will stay the same at the new scale. The formulation and the device must be understood as a coupled system, with each scale transition testing whether that relationship still holds.
Tangential flow filtration: the downstream step that most often breaks scale-up
Particle formation is only one of the two major physical challenges. Tangential flow filtration (TFF) is the standard downstream method for removing solvent, exchanging buffer, and concentrating LNPs. It passes the suspension across a semipermeable membrane under controlled transmembrane pressure, which lets smaller molecules through while keeping the particles. At every scale, its operating conditions need their own optimization and validation.
Development-scale TFF commonly uses small-volume hollow-fiber cartridges, whereas GMP production uses larger cartridges or parallel units. That transition can change both shear stress and effective membrane area. Transmembrane pressure, cross-flow velocity, number of diafiltration volumes, and membrane molecular weight cut-off each affect how well the process runs and the quality of the product. Even when mixing goes well, purification can still damage particles or cause them to vary.
Pallerla et al. reported a 100-fold scale increase in the SMNP program in Journal of Pharmaceutical Sciences in September 2025. Closed-system dilution and diafiltration supported the transition, while analytical characterization established consistent particle size, morphology, and polydispersity relative to preclinical batches. The case shows that downstream integrity can survive a large scale jump when the process is designed around it. For olfactory delivery approaches, including Lionbio’s nose-to-brain platform, TFF integrity similarly matters to particle uniformity and structure, on which potential bioavailability and nausea-reduction benefits depend.
Why batch synthesis reproducibility fails and what continuous manufacturing changes
Each run of batch synthesis starts with a fresh set of initial conditions. As batches grow, it gets harder to keep the formulation stable, the raw materials consistent, and the process under control. Each scale transition exposes those vulnerabilities again, making reproducibility a recurring problem. Understanding that reset explains the appeal of continuous manufacturing more clearly than a general claim that continuous processes are better.
Continuous manufacturing holds process conditions steady across a production run and supports real-time monitoring. With parallelization, it can make more while keeping the fluid dynamics that govern particle quality at small scale. Instead of enlarging a process and accepting a new mixing environment, the approach repeats controlled conditions. Its value lies in that continuity as well as throughput.
The practical limits are substantial, however. Continuous infrastructure costs a lot to set up, and the equipment already in use cannot always be adapted. Skilled staff are also needed to implement and maintain automation and process analytical technology. For complex nanoparticles, the materials lost, delays, and regulatory consequences when a GMP batch fails can make that spending worthwhile. The choice still depends on whether the program can build and sustain the control system that makes continuous operation useful.
How process analytical technology catches quality drift before it becomes a batch failure
Even a well-characterized process can drift during a run. Changes in FRR, shear, or membrane pressure may shift particle size, size distribution, or encapsulation efficiency without anything visible to the operator. Process analytical technology (PAT) supplies the missing visibility. It closes the feedback loop between operating conditions and product quality, making scale-up controllable rather than dependent on a satisfactory result at the end.
In-line or at-line dynamic light scattering can detect particle size changes during mixing or TFF before they become a batch failure. In-line fluorescence monitoring using intercalating dyes can also provide a real-time indicator of encapsulation efficiency during formulation. The FDA encouraged PAT adoption through its 2004 guidance, “PAT, A Framework for Innovative Pharmaceutical Development, Manufacturing, and Quality Assurance.” For LNPs, size distribution affects how well they work in clinical use and failed batches cost a lot, so monitoring is part of how manufacturing is set up.
Real-time indicators do not remove the need for complementary characterization. Pallerla et al. used CAD-HPLC for QS-21 and lipid quantification, dynamic light scattering for particle sizing, and TEM for morphological validation in the SMNP program. These orthogonal methods tested different aspects of product integrity at GMP scale. Together, they supported an assessment of equivalence with preclinical material that a single-method release test could not provide.
The regulatory framework that cGMP compliance imposes on nanoparticle manufacturers
These technical controls also have a legal role. Section 501(a)(2)(B) of the Federal Food, Drug, and Cosmetic Act requires drug products containing nanomaterials to be manufactured under current Good Manufacturing Practices. Because nanoparticles are sensitive to physical conditions, programs must understand the process to comply with the law. Producing the right particles once is not the same as establishing a reproducible manufacturing process.
Raw material variability makes that distinction concrete. Development work that uses just one lipid or polymer lot may miss variation that occurs when GMP production uses different source lots. Those differences can change particle properties even when equipment settings stay the same. Pallerla et al. identified managing raw material variability as a critical part of reproducibility in the SMNP program.
Safety and stability characterization must also address the material made by the qualified process. Hemolysis testing, stability studies under ICH-specified conditions, and in vitro safety assays must be performed on GMP-manufactured material. The manufacturing process is part of what is being qualified because it can alter the particles under evaluation. Readiness for clinical trials requires both technical success and the regulatory evidence to support it.
Biological challenges that scale-up cannot solve but cannot ignore: protein corona and formulation stability
GMP control does not make a particle’s biological behavior predictable from its behavior in buffer alone. On contact with physiological systems, proteins and other biomolecules adsorb to nanoparticle surfaces. The resulting corona can change particle size, surface charge, and access to targeting moieties. If those moieties become hidden, a particle designed for targeting can function more like a passive particle. Manufacturing consistency preserves the formulation, but it cannot by itself preserve a targeting advantage that biological exposure removes.
Surface charge presents a related design tension. Positively charged particles can enhance gene transfection and lysosomal escape, yet they can also disrupt cell membranes from the entry point to the target. Maintaining the development formulation’s charge profile at GMP scale is therefore part of preserving its safety and efficacy profile. A consistently manufactured surface must also remain appropriate in its intended environment.
Stability adds time to this problem. Clinical material must remain within specification through manufacture, release, shipping, and use, which is the window ICH stability studies characterize. Pallerla et al. found that SMNP material was stable under refrigeration, while frozen and elevated temperature conditions revealed degradation pathways relevant to clinical supply decisions. Formulation design must anticipate both biological exposure and storage conditions. Mucoadhesive coatings and stealth surfaces are consequently design choices to be considered before scale-up.
Why intranasal nanoparticle programs face a compounded version of the scale-up problem
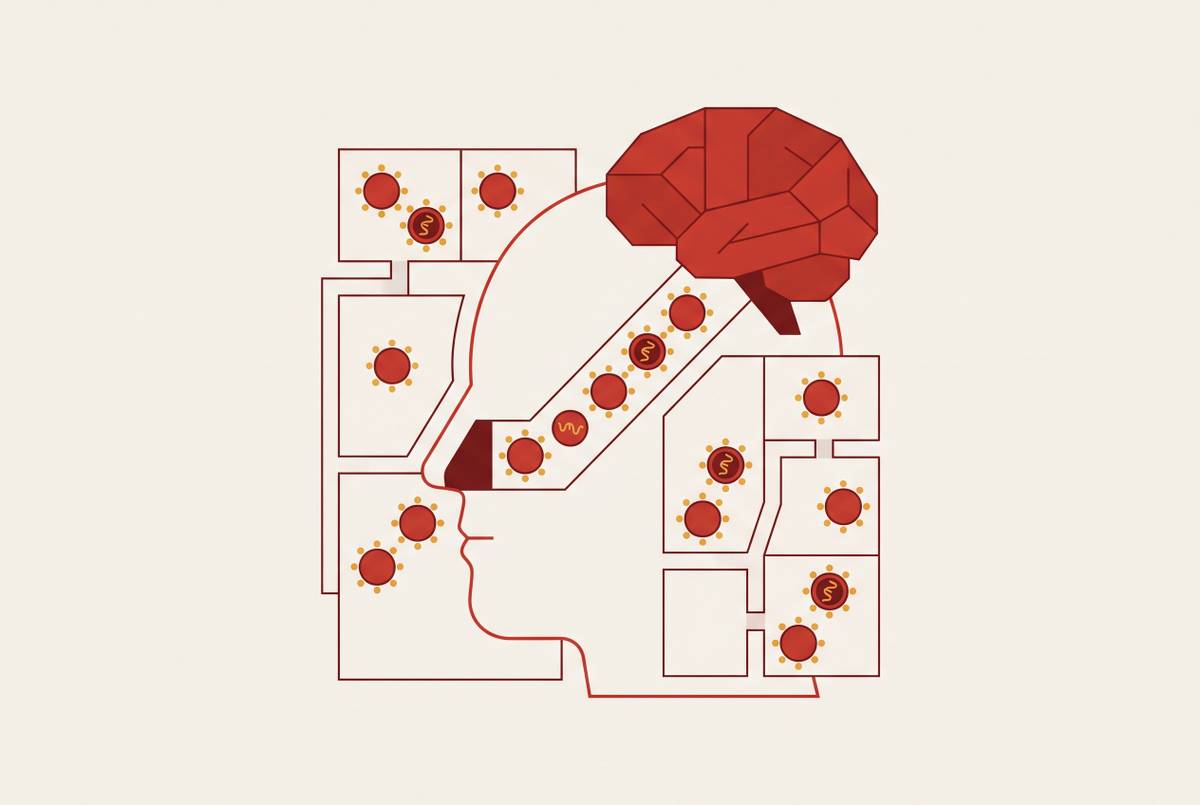
Particle quality must meet GMP requirements, while viscosity, mucoadhesion, and deposition fraction must support delivery to the olfactory epithelium at therapeutic levels. Enzymatic degradation, mucociliary clearance, and limited residence time on the nasal mucosa all constrain that delivery. Nanoparticle systems are engineered to address these barriers, but the engineering resides in formulation properties that must survive scale-up.
A study of a borneol-modified lipid nanoparticle nasal spray reported an olfactory deposition fraction of 23.58%, highlighting deposition as a measurable formulation outcome. The GMP formulation must retain the development viscosity profile because shifts can reduce the fraction of drug reaching the brain. Chitosan-based mucoadhesive coatings can extend nasal residence and improve brain targeting efficiency, but they also affect particle size, charge, and stability.
Lionbio is developing intranasal nanoparticle-based GLP-1 therapeutics on a platform built on more than 30 years of patented university nanoparticle delivery IP. Its approach illustrates the need to connect control of particle formation with control of delivery, from early formulation work through GMP manufacturing. For the peptide therapeutics it is developing, consistent FRR and mixing geometry are part of the mechanism for maintaining dose and bioavailability. Every decision about size, surface chemistry, or viscosity is therefore also a decision about potential clinical performance.
What a systematic scale-up program actually looks like from development batch to clinical trial material
A systematic program treats development-to-pilot and pilot-to-GMP transitions as distinct validation challenges. Each needs process characterization, analytical qualification, and stability work beyond a change in batch volume in the same proportion. FRR, total flow rate, TFF transmembrane pressure, and lipid concentration must be re-characterized at the new scale. Keeping the same formulation team involved helps keep the reasons for those controls clear at the next stage.
The SMNP case shows what that integration can achieve. Its closed-system process incorporated orthogonal characterization, ICH stability studies, and hemolysis assay benchmarking rather than adding them after manufacturing was complete. Pallerla et al. reported that cGMP-compliant material was ready for studies to support clinical trial authorization, with records that lay the groundwork for later commercial production. The achievement was not simply the scale increase, but the evidence that product integrity had survived it.
Continuous manufacturing with PAT offers an architecture for preserving that control through parallelization and real-time feedback. It does not replace the scientific work of identifying which variables may shift and designing experiments to test them. That work starts during formulation development because how the product is made is tied to its physical and biological design. Nanoparticle scale-up breaks when it is treated as a volume increase and succeeds when the program understands and controls the physics.
Sources
- Scale-up and cGMP manufacturing of next-generation vaccine adjuvant saponin/MPLA nanoParticles (SMNP) - ScienceDirect
- Progress on drug nanoparticle manufacturing: exploring the adaptability of batch bottom-up approaches to continuous manufacturing - ScienceDirect
- Scale-up and cGMP Manufacturing of Next-Generation Vaccine Adjuvant Saponin/MPLA NanoParticles (SMNP)
- Nanocarrier-based intranasal drug delivery for enhanced neurological disorders treatment